Aldehydes & ketones
The carbonyl group is polarized, with the carbon being electron-poor and the oxygen electron-rich, and most aldehyde and ketone chemistry follows from that. A nucleophile attacks the carbon, the π bond breaks, and the oxygen takes the electrons. What happens next depends on whether the resulting alkoxide can be protonated or can expel a leaving group.
The carbonyl groupPreparationOxygen nucleophilesCarbon nucleophilesNitrogen nucleophiles & reductionBaeyer–Villiger
01The carbonyl group
Structure, polarity and relative reactivity
The carbonyl carbon is sp² hybridized, so the group is planar with bond angles near 120°, and the π bond is formed from the overlap of carbon and oxygen p orbitals. Oxygen is considerably more electronegative than carbon, so the π electrons are pulled toward it; the oxygen carries a partial negative charge and the carbon a partial positive one. A second resonance structure with a full positive charge on carbon and a negative charge on oxygen may be drawn that highlights the polarization.
That polarization sets up two reactive sites. Nucleophiles attack the electron-poor carbon, and electrophiles, in practice, usually protons, coordinate to the oxygen lone pairs. Under basic conditions the nucleophile is strong enough to attack the neutral carbonyl directly. Under acidic conditions, protonating the oxygen makes the carbon far more electrophilic, which is why so many carbonyl additions are acid-catalyzed.
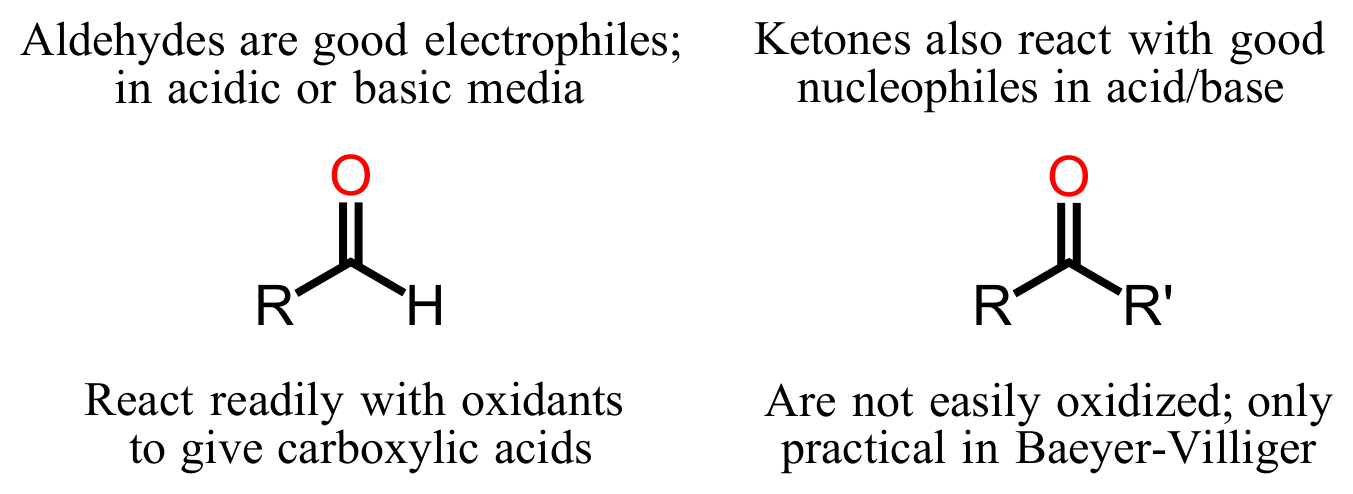
Aldehydes are generally more reactive than ketones, for two reasons. Sterically, an aldehyde has one alkyl group and one hydrogen at the carbonyl carbon rather than two alkyl groups, so less crowding of the approaching nucleophile. Electronically, an alkyl group donates electron density and reduces the partial positive charge, so two attached donating groups stabilize the carbonyl more than one does. The same argument explains why formaldehyde (no alkyl groups attached) is the most reactive of all.
02Preparation
Oxidation, ozonolysis and acylation
Oxidation of an alcohol is the most common route to carbonyl-containing functional groups. A secondary alcohol gives a ketone with any of the usual oxidants, since there is nowhere further to go due to the lack of another hydrogen on the alpha carbon. A primary alcohol is more versatile: chromic acid or KMnO₄ carries the oxidation past the aldehyde to the carboxylic acid, so a milder, selective reagent is needed to stop at the aldehyde. PCC or PDC in anhydrous dichloromethane serves that role.
Alkenes and alkynes provide two more routes. Ozonolysis cleaves a double bond and delivers the two carbonyl fragments, with a reductive workup such as zinc or dimethyl sulfide chosen so that any aldehyde formed survives. Hydration of an alkyne gives a ketone by Markovnikov addition of water, while the hydroboration–oxidation route on a terminal alkyne gives the aldehyde instead. Both of those alkyne additions are followed by tautomerism from an enol to the more stable carbonyl.
For an aryl ketone, Friedel–Crafts acylation is the standard method: an acid chloride with AlCl₃ attaches the acyl group directly to the ring, with the reaction stopping at one substitution since the introduced carbonyl is electron-withdrawing and thus deactivating. Acid chlorides also react with organocuprates to give ketones, and with a hindered hydride such as lithium tri(tert-butoxy)aluminium hydride to give aldehydes.
03Addition of oxygen nucleophiles
Hydrates, acetals and protecting groups
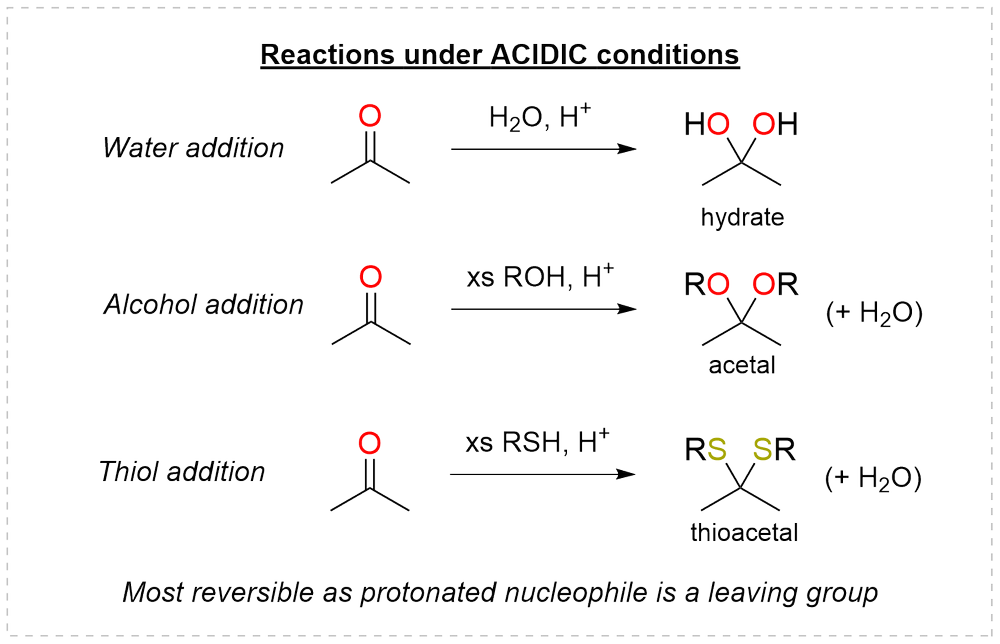
Water adds reversibly to a carbonyl to give a hydrate (a geminal diol), a carbon bearing two hydroxyl groups. For most ketones the equilibrium lies well to the left, so the hydrate is only a minor species; aldehydes give mixtures, and formaldehyde is almost completely hydrated in water. Electron-withdrawing groups next to the carbonyl in aldehydes or ketones push the equilibrium further toward the hydrate.
Alcohols also react with aldehydes and ketones but go one step further. One equivalent of alcohol gives the hemiacetal, which is usually not isolated. Under acidic catalysis, the hemiacetal hydroxyl is protonated, water leaves to give a resonance-stabilized oxocarbenium ion, and a second alcohol molecule attacks to give the acetal. Every step in the process is reversible, so the reaction is driven forward by removing water via distillation or a drying agent. The carbonyl may be regenerated by treating the acetal with aqueous acid.
That reversibility is what makes acetals so useful. An acetal is unreactive toward base and nucleophiles, so converting a ketone to an acetal protects it while basic chemistry is carried out elsewhere in the molecule. For example, reducing an ester, or adding a Grignard reagent to another site. Subsequently, mild aqueous acid will regenerate the carbonyl by hydrolyzing the acetal.
04Addition of carbon nucleophiles
Grignard, cyanide and the Wittig reaction
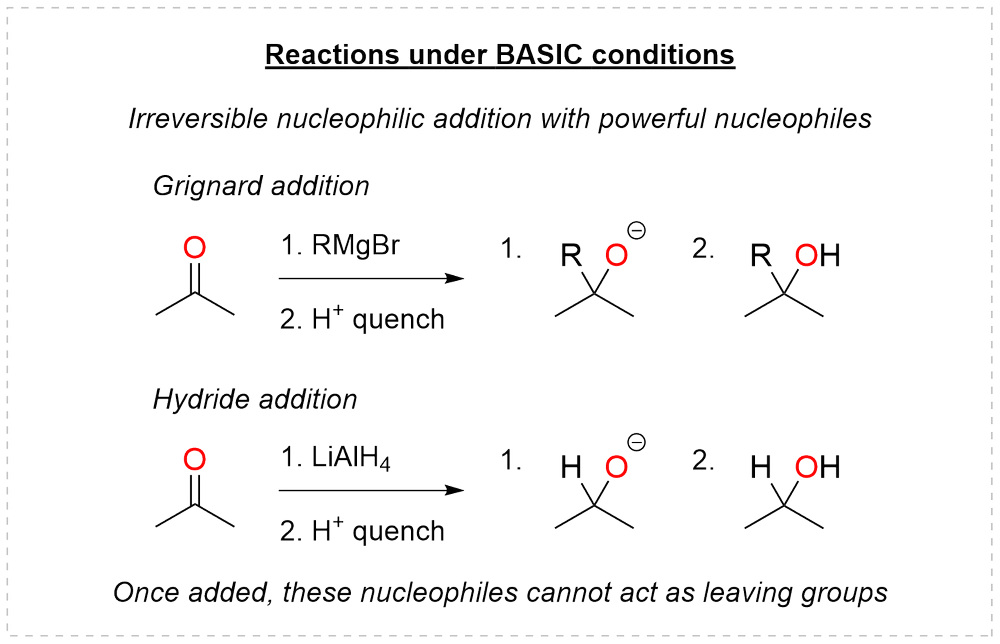
A Grignard reagent or an organolithium species attacks the carbonyl carbon irreversibly, and an aqueous workup protonates the resulting alkoxide. Formaldehyde gives a primary alcohol, any other aldehyde gives a secondary alcohol, and a ketone gives a tertiary alcohol. Working backwards from the alcohol to choose the carbonyl and the organometallic partner is a standard retrosynthetic step.
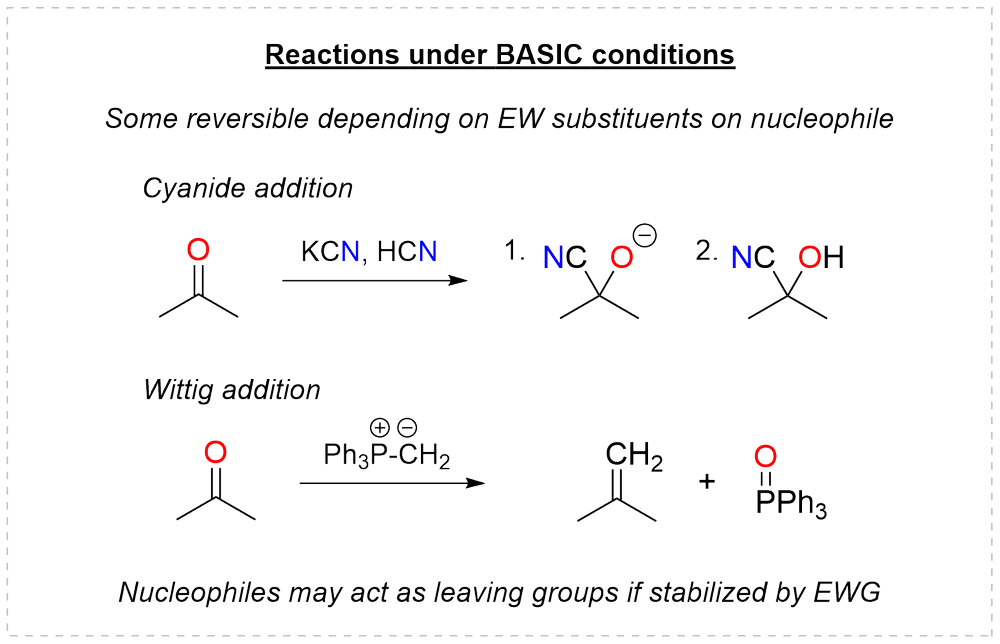
Cyanide adds reversibly to give a cyanohydrin, a molecule with both a hydroxyl and a nitrile on the same carbon. Hydrolysis converts the nitrile to a carboxylic acid, and reduction converts it to a primary amine, which makes cyanohydrins quite versatile intermediates.
The Wittig reaction is slightly different in that the oxygen of the carbonyl is not found in the addition product. A phosphonium ylide adds to the carbonyl, the alkoxide bonds with the positive phosphorus atom to give a four-membered oxaphosphetane, and that ring collapses to expel triphenylphosphine oxide and form a C=C bond. The advantage over an elimination route to alkenes is that the double bond appears exactly where the carbonyl carbon was, with no rearrangement possible.
05Nitrogen nucleophiles and reduction
Imines, enamines and hydride reagents
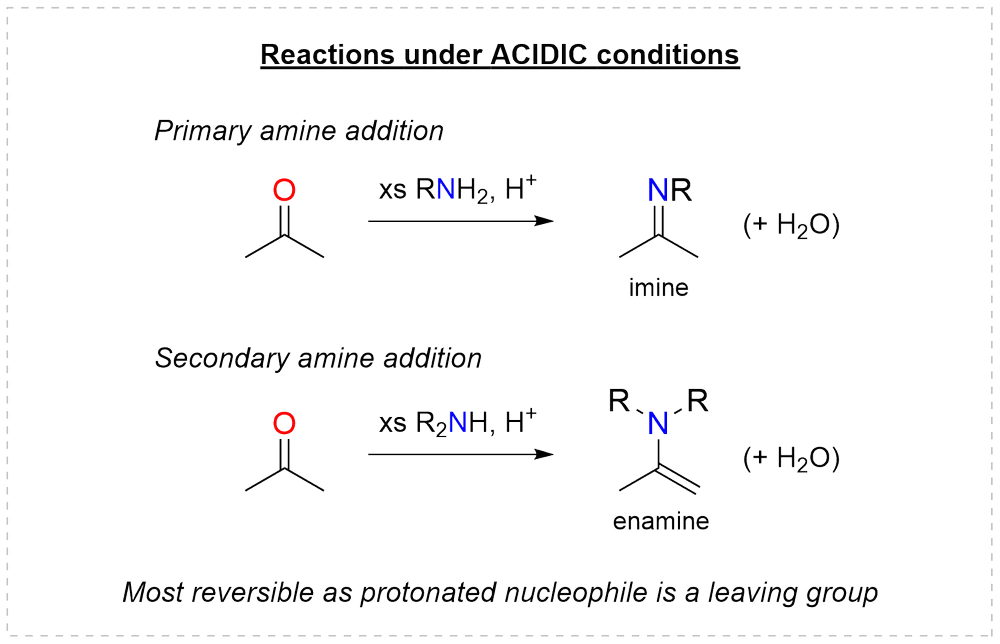
A primary amine adds to the carbonyl of an aldehyde or ketone and the tetrahedral intermediate then loses water to give an imine, with the C=N double bond directly replacing C=O. The reaction rate is greatest around pH 4–5, which is an experimental compromise. Acid is needed to protonate the hydroxyl so water can leave, but too much acid protonates the amine and quenches the nucleophile. A secondary amine reacts in a similar way, however there is no extra proton to lose from nitrogen, so it eliminates from the α-carbon instead and gives an enamine. If the elimination is possible from two different alpha carbons, the more substituted alkene is favoured.
Reduction adds hydride to the carbonyl carbon to give an alcohol; an aldehyde gives a primary alcohol, and a ketone a secondary one. Sodium borohydride is mild enough to use in alcohol or water as solvent and reduces aldehydes and ketones while leaving esters and carboxylic acids intact. Lithium aluminium hydride is far more reactive, and reduces essentially every carbonyl including esters and acids. It also reacts violently with water, so it is used in dry ether or THF with a separate aqueous workup.
Other reductions remove the oxygen from the carbonyl altogether. Clemmensen reduction, with Zn amalgam in HCl, and Wolff–Kishner reduction, via the hydrazone with strong base and heat. Both convert C=O to CH₂. These reactions apply to Friedel–Crafts chemistry; acylate the ring, then reduce, and an alkylbenzene results, which could be problematic via direct alkylation. Choosing between these methods depends on what else is present in the molecule at hand since one protocol needs strong acid, the other strong base.
06Baeyer–Villiger oxidation
Inserting an oxygen next to the carbonyl
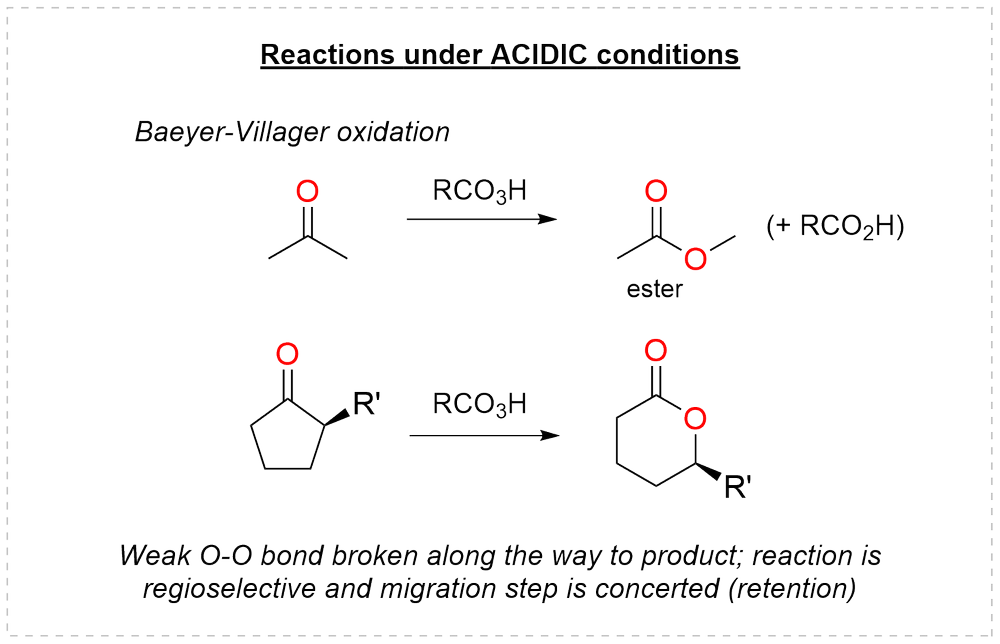
Ketones resist most oxidants, since there is no hydrogen on the carbonyl carbon to remove, but a peroxyacid converts ketones to esters. The peroxyacid adds to the carbonyl to give a tetrahedral Criegee intermediate, and the weak O–O bond then breaks as one of the two groups on the carbonyl carbon migrates onto the adjacent oxygen; a carboxylic acid leaves as the by-product. With a cyclic ketone the same insertion enlarges the ring by one atom and the product is a lactone.
Two features make the outcome predictable. The reaction is regioselective; the more electron-rich group that is better able to support positive charge migrates, so the order runs tertiary > secondary > primary > methyl, and with an unsymmetrical ketone we can predict which side the oxygen is inserted on. Also, because the migration step is known to be concerted, the migrating carbon does not reorganize, so a stereocentre at that position is retained with no loss of configuration.
Reaction summary
Each transformation is listed here. Read it right to left when planning a synthesis.
| Substrate | Reagents | Product | What controls it |
|---|---|---|---|
| 1° alcohol | PCC or PDC, CH₂Cl₂ | Aldehyde | Mild and anhydrous, so oxidation stops at the aldehyde |
| 2° alcohol | H₂CrO₄ or PCC | Ketone | No further oxidation is possible; reagent choice is flexible |
| Alkene | 1. O₃ 2. Zn or (CH₃)₂S | Two carbonyls | Reductive workup preserves any aldehyde formed |
| Arene | RCOCl, AlCl₃ | Aryl ketone | Acylium ion cannot rearrange; stops at one substitution |
| Aldehyde or ketone | H₂O, trace acid or base | Hydrate | Reversible and usually unfavourable; EWGs shift equilibrium to the right |
| Aldehyde or ketone | 2 ROH or a diol, H⁺, −H₂O | Acetal | Driven by removing water; reversed by adding aqueous acid |
| Aldehyde or ketone | 1. RMgX 2. H₃O⁺ | 2° or 3° alcohol | Irreversible C–C bond formation; the workup protonates the alkoxide formed |
| Aldehyde or ketone | HCN or NaCN/H⁺ | Cyanohydrin | Reversible; the nitrile converts to an acid or an amine |
| Aldehyde or ketone | Ph₃P=CR₂ | Alkene | Wittig; C=C forms where C=O was, with no rearrangement |
| Aldehyde or ketone | RNH₂, pH 4–5 | Imine | Mildly acidic pH balances leaving-group need against amine basicity |
| Aldehyde or ketone | R₂NH, H⁺ | Enamine | No N–H left, so elimination occurs from the α-carbon |
| Aldehyde or ketone | NaBH₄, CH₃OH | 1° or 2° alcohol | Mild; leaves esters and carboxylic acids untouched |
| Ketone | Zn(Hg), HCl | Alkane | Clemmensen; acidic conditions, C=O to CH₂ |
| Ketone | 1. H₂NNH₂ 2. KOH, heat | Alkane | Wolff–Kishner; the basic alternative to Clemmensen |
| Ketone | RCO₃H | Ester | Baeyer–Villiger; the better cation-stabilizing group migrates |
| Cyclic ketone | RCO₃H | Lactone | Ring expands by one oxygen; migration is concerted, so configuration is retained |
Six questions before you move on
Try to work out an answer on paper, then reveal to check. If your reasoning is right but the answer is wrong, you are closer than you think.
Q1Why is an aldehyde more reactive toward nucleophiles than a ketone?
Two reasons that reinforce each other. Sterically, one alkyl group and one hydrogen leave the carbonyl carbon more open to attack than two alkyl groups. Electronically, alkyl groups donate electron density and reduce the partial positive charge, so a ketone’s carbonyl carbon is less electrophilic.
Q2You need to make the aldehyde from 1-butanol. Why will chromic acid not work, and what will?
Chromic acid oxidizes a primary alcohol straight through the aldehyde to butanoic acid, because the aldehyde hydrates in the aqueous medium and is oxidized further. PCC or PDC in dichloromethane is mild and anhydrous and stops at butanal.
Q3Acetal formation is reversible at every step. How do you drive it forward, and how do you reverse it?
To drive forward, remove the water as it is formed, by distillation or by adding a drying agent, and use excess alcohol. To reverse the reaction, add water back. Aqueous acid hydrolyzes the acetal to the carbonyl and two alcohols. Le Châtelier’s principle applies in both directions.
Q4A molecule contains both a ketone and an ester, and you need to reduce only the ester. What do you do?
Protect the ketone as its cyclic acetal with ethylene glycol and acid, reduce the ester with LiAlH₄, then hydrolyze the acetal with aqueous acid. The acetal is inert to hydride, and reagent selectivity alone will not help here, since LiAlH₄ would reduce the ketone too.
Q5Imine formation is fastest near pH 4–5 and slow in both strong acid and neutral base. Explain this difference.
Losing water from the tetrahedral intermediate needs acid to protonate the hydroxyl, but strong acid protonates the amine and destroys the nucleophile. Mildly acidic conditions leave enough free amine to attack while still assisting the dehydration.
Q6Why is a Wittig reaction preferred over dehydrating an alcohol when you need an alkene in a specific position?
The Wittig places the C=C exactly where the C=O was, and since no carbocation is involved, nothing can rearrange or eliminate in a competing direction. Acid-catalyzed dehydration goes through a cation, which may rearrange, and Zaitsev’s rule may put the double bond somewhere you did not want it.